Non-compliance with GMP in the pharmaceutical sector often arises from systemic weaknesses, including data integrity issues, inadequate investigations, and ineffective corrective actions. Identifying and addressing these vulnerabilities is essential for maintaining a robust quality assurance framework.
In the pharmaceutical sector, adherence to Good Manufacturing Practices (GMP) is paramount for ensuring product safety and efficacy. However, non-compliance can often stem from a confluence of systemic weaknesses rather than isolated errors. Identifying these vulnerabilities is crucial for maintaining a robust quality assurance framework.
1. Issues with Data Integrity
Data integrity remains a critical challenge, as any lapse can compromise the quality control processes that rely on accurate information. Problems can manifest in various forms, including incomplete records, backdated entries, and poorly managed electronic systems. For example, the FDA has previously cited companies for discrepancies in their data management practices, highlighting the severe implications of inadequate data governance.
Quality assurance must go beyond merely rectifying data issues. It requires a comprehensive evaluation of the problem's scope, the affected systems, potential impacts on released products, and whether there are additional processes that could yield similar discrepancies. The focus should be on rectifying the underlying quality system rather than just fixing isolated records.
2. Inadequate Investigative Processes
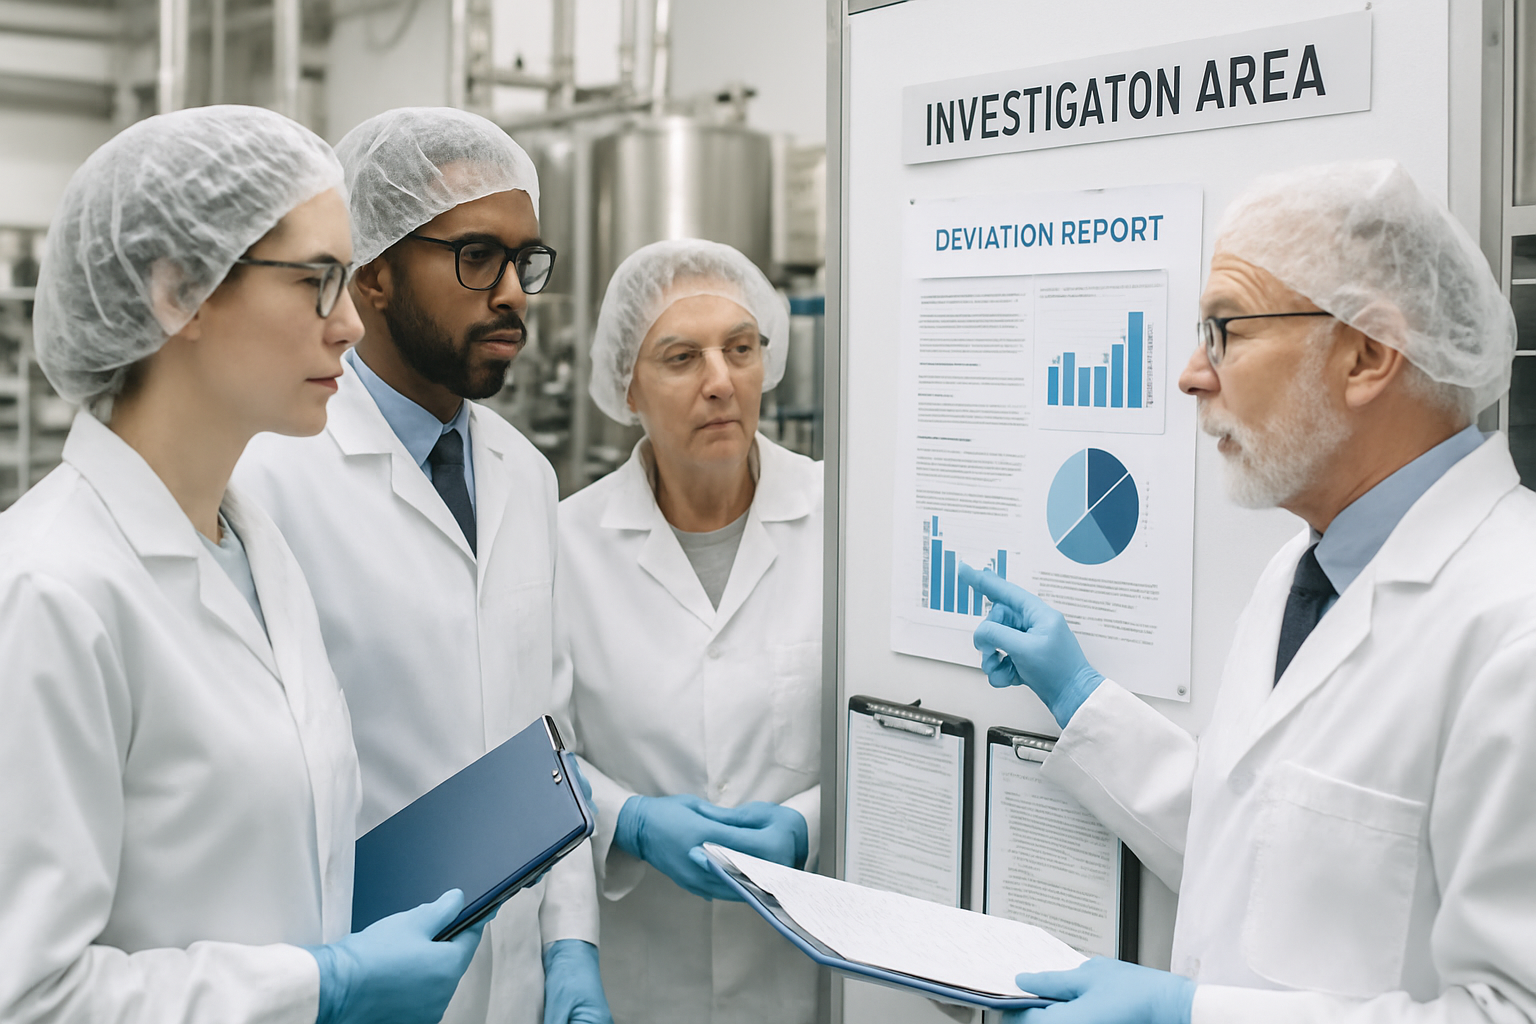
Another common pitfall is the superficial nature of investigations into deviations and out-of-specification (OOS) results. Often, companies may merely attribute issues to operator error without delving into the root causes. This limited approach fails to address whether the error is part of a broader systemic issue.
Effective investigations should encompass a thorough analysis of the incident, assess its overall impact, and establish scientifically valid reasons for the failure. It is essential that OOS results are not dismissed as mere analytical errors without comprehensive investigations that consider all relevant factors, including manufacturing processes.
3. Ineffective Corrective and Preventive Actions
Loading flowchart…
Corrective and Preventive Actions (CAPA) must be implemented effectively to address root causes rather than simply amending procedures or retraining staff. For instance, if repeated deviations arise from batch record errors, the focus should shift to identifying whether the issues stem from poor documentation practices or inadequate supervision.
According to ICH Q10, a successful pharmaceutical quality system should foster continuous improvement and accountability. Thus, the effectiveness of CAPA should be validated through objective evidence, not merely by confirming that an action has been taken.
4. Insufficient Quality Unit Authority
The role of the quality unit is critical, yet it often falls short due to limited authority and resources. An effective quality unit must have the independence to oversee processes without being compromised by commercial interests. Regulatory bodies have highlighted situations where quality units failed to enforce compliance, underscoring the need for robust oversight mechanisms.
Quality assurance teams should possess the authority to halt production, escalate issues, and conduct thorough investigations when necessary. A strong quality culture is reflected in the decisions made during critical moments.
5. Non-Adherence to Established Procedures
Merely having a Standard Operating Procedure (SOP) does not guarantee compliance. Discrepancies often arise between documented procedures and actual practices due to outdated procedures, insufficient training, or procedural complexity.
To mitigate risks, organizations must rigorously compare approved procedures with on-the-ground practices and documentation. Systematic breaches should prompt a review of the procedures themselves to ensure they remain relevant and practical.
6. Ineffective Manufacturing Process Control
The validation of manufacturing processes does not conclude once initial qualifications are achieved. Continuous monitoring is essential to ensure processes remain capable and under control. Indicators such as high variability in production yield or recurrent equipment failures can signal underlying control issues.
The FDA advocates for a continual process verification approach that monitors trends rather than waiting for batch failures. A proactive stance is vital for maintaining process integrity.
7. Mishandling of OOS, Out-of-Trend, and Deviations
Prematurely attributing negative results to laboratory errors without sufficient investigation is a dangerous practice. A thorough examination should include not only laboratory conditions but also consider manufacturing practices and sampling methods.
This comprehensive approach is essential, as deviations—even those within specifications—can indicate significant underlying trends. A well-trained quality assurance team should analyse historical data, process capabilities, and previous analytical trends to inform their assessments.
8. Challenges with Documentation and Record Management
The integrity of documentation is as critical as product quality. Inadequate record-keeping, such as missing forms or improperly maintained notebooks, can obscure quality control assessments. The FDA has documented instances where personnel have used unauthorized records, raising substantial concerns about compliance.
A practical test of documentation efficacy is whether an independent reviewer can reconstruct events based solely on original GMP documents. If this is not achievable, the documentation system requires urgent evaluation.
Identifying Systemic Compliance Weaknesses
Organizations should not wait for regulatory inspections to discover compliance issues. Proactive internal audits and management reviews can reveal systemic weaknesses, including:
- Recurring deviations from similar root causes
- Repeated OOS results
- Consistently extended or reopened CAPAs
- Inconsistencies in audit trails
- Frequent equipment breakdowns
- Unexplained trends in processes
- Insufficient evidence in quality reviews
Recognizing the significance of these observations from a systems perspective is crucial. Addressing issues as independent occurrences can mask larger, systemic problems that need resolution.
Focus Areas for Regulatory Inspectors
During inspections, regulators assess not only the nature of issues but also the efficacy of the quality system in identifying and addressing them. Key areas of scrutiny include the depth of investigations, product impact assessments, historical data analysis, and the effectiveness of CAPA measures.
Responses to inspection findings should address systemic issues rather than superficial symptoms. According to ICH Q9, quality risk management should be grounded in scientific principles. Serious GMP compliance failures tend to be systemic, often intertwined with data integrity, ineffective investigations, and inadequate documentation processes. Building a resilient quality system will require companies to focus on establishing a reliable recording system and conducting thorough investigations based on solid evidence.
Comments (0)
Loading comments…
Checking sign-in status…